McDermott and Bull executive Search
 

Other News

CAMZYOS is the only FDA-approved therapy for the treatment of oHCM in a pediatric population Approval based on positive results of Phase 3 SCOUT-HCM trial, building on the extensive clinical evidence for CAMZYOS in adults with oHCM

PRINCETON, N.J.–(BUSINESS WIRE)– Bristol Myers Squibb (NYSE: BMY) today announced that the U.S. Food and Drug Administration (FDA) approved CAMZYOS® (mavacamten) for the treatment of symptomatic obstructive hypertrophic cardiomyopathy (oHCM) to improve functional capacity and symptoms in adults and pediatric patients weighing 30 kg (66 lbs) or more. CAMZYOS now […]

BioCardia Files Revised De Novo Pre-Submission for FDA Approval of Helix™ Catheter

SUNNYVALE, Calif., Oct. 01, 2026 (GLOBE NEWSWIRE) — BioCardia®, Inc. [Nasdaq: BCDA], a global leader in cellular and cell-derived therapeutics for the treatment of cardiovascular and pulmonary diseases, today reported it has completed its revised Pre-Submission to FDA Center for Devices and Radiological Health (CDRH) under its Q-Submission program for the approval of its Helix Catheter (“Helix”) for therapeutic and diagnostic agent delivery to the heart. In May 2026, the Company and FDA met and reviewed our initial DeNovo pre-submission. FDA agreed that there are two pathways for Helix marketing clearance and raised no concerns regarding the Helix safety data, device performance, or compatibility with general classes of agents. In August 2026, the FDA confirmed and accepted the meeting minutes as summarized by BioCardia. The revised De Novo pre-submission for the Helix catheter incorporates the advice from FDA CDRH detailed in the minutes. Should FDA enable Helix to be the first device of its kind cleared for market release by FDA, the extensive clinical data and safety profile from 15 clinical trials including nearly 500 patients presents opportunities for FDA to ensure that follow-on catheters demonstrate non-inferiority to the Helix catheter safety profile and clinical experience prior to their marketing clearance. The FDA targets providing an initial acceptance/refusal review within 15 calendar days and aims to issue written feedback or hold a meeting within 70 calendar days. An independent FDA clearance of Helix has the potential to expand BioCardia’s opportunities to partner with developers of investigational cell, gene and protein therapeutics requiring targeted delivery to the heart. About BioCardia BioCardia, Inc., headquartered in Sunnyvale, California, is a global leader in cellular and cell-derived therapeutics for the treatment of cardiovascular and pulmonary disease. CardiAMP™ autologous and CardiALLO™ allogeneic cell therapies are the Company’s biotherapeutic platforms with three cardiac clinical stage product candidates in development. These therapies are enabled by its Helix biotherapeutic delivery and Morph® vascular navigation product platforms, and soon the Heart3D™ fusion imaging platform. BioCardia selectively partners on biotherapeutic delivery with peers developing important biologic therapies. For more information, visit www.biocardia.com. Forward-Looking Statements This press release contains forward-looking statements that are subject to many risks and uncertainties. Forward-looking statements include, among other things, statements relating to the timing and potential outcome of BioCardia’s regulatory submissions for the Helix catheter. These forward-looking statements are made as of the date of this press release. We may use terms such as “believes,” “estimates,” “anticipates,” “expects,” “plans,” “intends,” “may,” “could,” “might,” “will,” “should,” “approximately” or other words that convey the uncertainty of future events or outcomes to identify these forward-looking statements. Although we believe that we have a reasonable basis for each forward-looking statement contained herein, we caution you that forward-looking statements are not guarantees of future performance and that our actual results may differ materially from the forward-looking statements contained in this press release. Factors that could cause or contribute to such differences include, but are not limited to, the Company’s liquidity position and its ability to raise additional funds, as well as the Company’s ability to successfully progress its clinical trials and regulatory programs. Additional factors that could materially affect actual results can be found in BioCardia’s Form 10-K filed with the Securities and Exchange Commission on March 24, 2026, under the caption titled “Risk Factors” and in its subsequently filed Quarterly Reports on Form 10-Q. BioCardia expressly disclaims any intent or obligation to update these forward-looking statements, except as required by law. Media Contact: Miranda Peto, Investor RelationsEmail: mpeto@BioCardia.comPhone: 650-226-0120 Investor Contact: David McClung, Chief Financial OfficerEmail: investors@BioCardia.comPhone: 650-226-0120

Balt Announces First Patients Enrolled in Silk Vista Baby™ PIONEER Study

IRVINE, Calif., Oct. 01, 2026 (GLOBE NEWSWIRE) — Balt announced today it has enrolled the first patients in North America in the premarket approval (PMA) clinical trial, PIONEER (PIvotal Study Of the Silk Vista Baby Flow Diverters iN the TrEatmEnt of Unruptured Intracranial AneuRysms). Pioneer is the first prospective trial exclusively assessing the role of flow diversion in the treatment of aneurysms in distal small arteries in the brain. This landmark trial is actively enrolling at hospitals in both the U.S. and Canada and is the U.S. approval study for the Silk Vista Baby flow diverter. Co-Primary Investigator, Vitor Mendes-Pereira, M.D., MSc (St. Michael’s Hospital, Toronto, Canada) stated, “The Silk Vista Baby has been a paradigm shift in the care of neurovascular patients. It provides physicians with a significant improvement on access to distal aneurysms with a low profile microcatheter over the treatment options we have available and enables us to provide better care for more patients.” The Silk Vista Baby device was the first flow diverter in the world that delivers through a small 0.017-inch diameter microcatheter. Obtaining European Union regulatory approval in 2018, the device has been used to treat thousands of aneurysm patients around the world and studied extensively in several clinical studies. Patients with distal brain aneurysms often have those aneurysms arise off very small blood vessels and the endovascular treatment options often require access devices that are too big, or the anatomy is not suitable for open surgical solutions. “The PIONEER study is an important advance for the care of aneurysm patients in the U.S.,” said co-Primary Investigator, Ricardo Hanel, M.D., PhD (Baptist Medical Center, Jacksonville, FL), “This study will add tremendously to our knowledge of how to care for patients with distal aneurysms and potentially brings another life-saving tool to the patients of the United States. One that has already changed care for much of the rest of the world.” Balt CEO Pascal Girin commented, “PIONEER marks the second PMA trial for Balt, following the groundbreaking STEM study which resulted in the PMA approval of the SquidTM Liquid Embolic for chronic subdural hematoma patients. We are currently working on multiple additional PMA studies and are committed to continue bringing Balt’s innovative solutions to novel indications for physicians and patients around the world.” About Balt: Since it was established in 1977, Balt has worked with interventional physicians to develop devices to treat complex life-threatening neurovascular diseases such as ischemic strokes, aneurysms and other hemorrhagic conditions including arteriovenous malformations, dural arteriovenous fistulas and chronic subdural hematomas. A pioneer in the neurovascular field, Balt designs, manufactures and distributes the broadest portfolio of products in the neurovascular space and is now focused on expanding its geographic presence in the U.S. Visit Balt at https://www.baltgroup.com/   ContactGreg ChodaczekGilmartin Groupgreg@gilmartinir.com

Orchestra BioMed Exceeds Full Enrollment Target in the BACKBEAT Global Pivotal Trial of AVIM Therapy

330 patients randomized as of September 30, 2026, surpassing full trial enrollment targetCompany reiterates prior guidance: primary endpoint data to be submitted for late-breaking presentation consideration at a major cardiovascular conference in Q2 2027 Data expected to support future marketing application submissions by Medtronic to the FDA and other global regulatory agencies NEW HOPE, Pa., Oct. 01, 2026 (GLOBE NEWSWIRE) — Orchestra BioMed Holdings, Inc. (Nasdaq: OBIO) (“Orchestra BioMed” or the “Company”), a biomedical company accelerating high-impact technologies to patients through strategic partnerships with market-leading medical device companies, today announced that it has surpassed full enrollment in the BACKBEAT Global Pivotal Trial (“BACKBEAT Trial”), exceeding its previously announced target of 316 randomized patients. The BACKBEAT Trial is evaluating Atrioventricular Interval Modulation Therapy (“AVIM Therapy”) in pacemaker-indicated patients with uncontrolled hypertension despite the use of medication. Hypertension is the most common modifiable risk factor for death worldwide and the most common comorbidity among the pacemaker population.1 The BACKBEAT Trial (NCT06059638) is being conducted with Medtronic plc (“Medtronic”), the Company’s strategic collaborator for the BACKBEAT Trial and, subject to regulatory approval, for the commercialization of AVIM Therapy in pacemaker-indicated patients with uncontrolled hypertension despite the use of medication. As of September 30, 2026, 330 patients have been randomized in the BACKBEAT Trial at 81 sites across 13 countries.The BACKBEAT Trial’s protocol, approved by the U.S. Food and Drug Administration (the “FDA”), specifies 284 evaluable randomized subjects for the primary endpoint analysis, with a total enrollment target of 316 patients to account for potential loss to follow-up. The BACKBEAT Trial is a global, multi-center, prospective, randomized, double-blind pivotal trial. Patients are randomized 1:1 to AVIM Therapy combined with continued medical therapy and pacing (treatment) or continued medical therapy and standard pacing alone (control). The primary efficacy endpoint of the BACKBEAT Trial is the between-group difference in the change in 24-hour ambulatory systolic blood pressure (“aSBP”) at three-month follow-up from pre-randomization baseline. The primary safety endpoint is freedom from unanticipated serious adverse device events in the AVIM Therapy arm at three-month follow-up. The protocol-specified sample size provides greater than 90% statistical power for both primary endpoints and is designed to detect a between-group difference of at least 5 mmHg in aSBP. David Hochman, Chairman & Chief Executive Officer of Orchestra BioMed, commented, “Achieving and surpassing full enrollment of the BACKBEAT Trial is a defining milestone for Orchestra BioMed, for our collaboration with Medtronic and potentially for the millions of high cardiovascular risk patients who remain above blood pressure target despite the medications available to them today. We are grateful for and excited about the enrollment momentum built across our global trial site network over the last year. I want to thank the patients and their families who chose to participate, and the clinical teams and investigators at our trial sites globally whose commitment made this milestone possible. Our focus now turns to completing three-month follow-up, database lock, and the delivery of a rigorous, well-powered data set next spring.” Robert C. Kowal, M.D., Ph.D., Vice President and General Manager for Cardiac Pacing Therapies in the Medtronic Electrophysiology Therapies operating unit, stated, “For decades, cardiac pacing has been foundational in the management of rhythm disorders. Reaching full enrollment in the BACKBEAT Trial brings us a big step closer to understanding whether AVIM Therapy can extend the role of cardiac pacing therapy to include hypertension management for patients at increased cardiovascular risk. We are proud of the collaborative work our team and Orchestra BioMed’s team have done alongside investigators around the world, and we look forward to the primary endpoint results.” Vivek Reddy, M.D., Executive Chairman of the BACKBEAT Clinical Steering Committee, Director of Cardiac Arrhythmia Services at The Mount Sinai Fuster Heart Hospital and Director of Electrophysiology for the Mount Sinai Health System, commented, “Randomized, double-blind evidence is the highest standard by which a new therapeutic modality earns its place in clinical practice, and that is exactly what the BACKBEAT Trial was designed to generate. The pilot data for AVIM Therapy have been exciting, consistent and mechanistically coherent, but the field needs a pivotal trial to answer the question definitively. On behalf of the Steering Committee, I want to recognize the investigators, coordinators and other professionals whose work enabled the trial to reach full enrollment, and I look forward to seeing the three-month primary endpoint results.” Clinical, Regulatory and Commercial Next Steps Orchestra BioMed and Medtronic intend to submit the BACKBEAT Trial’s primary endpoint results for consideration as a late-breaking clinical trial presentation at a major cardiovascular conference in the second quarter of 2027. Assuming positive results from the BACKBEAT Trial, Medtronic plans to submit AVIM Therapy marketing applications to the FDA and other global regulatory agencies promptly. About Orchestra BioMed Orchestra BioMed is a biomedical innovation company accelerating high-impact technologies to patients through strategic collaborations with market-leading global medical device companies. The Company’s two flagship product candidates, Atrioventricular Interval Modulation (AVIM) Therapy and Virtue Sirolimus AngioInfusion Balloon (Virtue SAB), are currently undergoing pivotal clinical trials for their lead indications, each representing multi-billion-dollar annual global market opportunities. AVIM Therapy is a bioelectronic treatment for hypertension, the leading risk factor for death worldwide, and is designed to be delivered by a pacemaker and achieve immediate, substantial and sustained reductions in blood pressure in patients with hypertensive heart disease. The Company has a strategic collaboration with Medtronic, one of the largest medical device companies in the world and the global leader in cardiac pacing therapies, for the development and commercialization of AVIM Therapy for the treatment of uncontrolled hypertension in pacemaker-indicated patients. AVIM Therapy has FDA Breakthrough Device Designations for these patients, as well as an estimated 7.7 million total patients in the U.S. with uncontrolled hypertension despite medical therapy and increased cardiovascular risk. Virtue SAB is a highly differentiated, first-of-its-kind non-coated drug delivery angioplasty balloon system designed to deliver a large liquid dose of proprietary extended-release formulation of sirolimus, SirolimusEFR™, for the treatment of atherosclerotic artery disease, the leading cause of mortality worldwide. Virtue SAB has been granted Breakthrough Device Designation by the FDA for the treatment of coronary in-stent restenosis, coronary small vessel disease and below-the-knee peripheral artery disease. For further information about Orchestra BioMed, please visit www.orchestrabiomed.com, and follow us on LinkedIn. About AVIM Therapy AVIM Therapy is an investigational therapy compatible with standard dual-chamber pacemakers designed to substantially and persistently lower blood pressure. It has been evaluated in pilot studies in patients with hypertension who are also indicated for a pacemaker. In MODERATO II, a double-blind, randomized pilot study, AVIM Therapy-treated patients had a mean reduction of 11.1 mmHg in 24-hour ambulatory systolic blood pressure (aSBP) from pre-randomization baseline at six months, representing a net reduction of 8.1 mmHg versus control patients. At the same time point, the net reduction in office systolic blood pressure (oSBP) was 12.3 mmHg versus control. Additional MODERATO II data showed an immediate mean oSBP reduction of 13.2 mmHg when AVIM Therapy was activated during setup before randomization, with 97% of patients achieving a reduction of at least 5 mmHg. In addition to reducing blood pressure, clinical results using AVIM Therapy demonstrate improvements in cardiac function and hemodynamics. The BACKBEAT (BradycArdia paCemaKer with atrioventricular interval modulation for Blood prEssure treAtmenT) global pivotal trial is evaluating the safety and efficacy of AVIM Therapy in lowering blood pressure in patients who have systolic blood pressure above target despite anti-hypertensive medication and who are indicated for or have recently received a dual-chamber cardiac pacemaker. AVIM Therapy has been granted two Breakthrough Device Designations by the FDA for the treatment of uncontrolled hypertension in patients who have increased cardiovascular risk. Forward-Looking Statements Certain statements included in this press release that are not historical facts are forward-looking statements for purposes of the safe harbor provisions under the United States Private Securities Litigation Reform Act of 1995. Forward-looking statements generally are accompanied by words such as “believe,” “may,” “will,” “estimate,” “continue,” “anticipate,” “intend,” “expect,” “should,” “would,” “plan,” “predict,” “potential,” “seem,” “seek,” “future,” “outlook” and similar expressions that predict or indicate future events or trends or that are not statements of historical matters. These forward-looking statements include, but are not limited to, statements relating to the timing, implementation, results and design of the Company’s ongoing pivotal trials, the timing of regulatory submissions, realizing the clinical and commercial value of the Company’s product candidates, the potential safety and efficacy of the Company’s product candidates, the size of the potential markets for the Company’s product candidates, and the ability of the Company’s partnerships to accelerate clinical development. These statements are based on various assumptions, whether or not identified in this press release, and on the current expectations of the Company’s management and are not predictions of actual performance. These forward-looking statements are provided for illustrative purposes only and are not intended to serve as and must not be relied on as a guarantee, an assurance, a prediction, or a definitive statement of fact or probability. Actual events and circumstances are difficult or impossible to predict and may differ from assumptions. Many actual events and circumstances are beyond the control of the Company. These forward-looking statements are subject to a number of risks and uncertainties, including changes in domestic and foreign business, market, financial, political, and legal conditions; risks related to regulatory approval of the Company’s commercial product candidates and ongoing regulation of the Company’s product candidates, if approved; the timing of, and the Company’s ability to achieve expected regulatory and business milestones; the impact of competitive products and product candidates; and the risk factors discussed under the heading “Item 1A. Risk Factors” in the Company’s Annual Report on Form 10-K for the year ended December 31, 2025, which was filed with the SEC on March 12, 2026. The Company operates in a very competitive and rapidly changing environment. New risks emerge from time to time. Given these risks and uncertainties, the Company cautions against placing undue reliance on these forward-looking statements, which only speak as of the date of this press release. The Company does not plan and undertakes no obligation to update any of the forward-looking statements made herein, except as required by law. Investor Contact:Silas NewcombOrchestra BioMedsnewcomb@orchestrabiomed.com Media Contact:Nina PremuticoOrchestra BioMednpremutico@orchestrabiomed.com 1 World Health Organization, Global Report on Hypertension 2025 (an estimated 1.4B adults aged 30–79 with hypertension in 2024, 33% of that age group; ~23% of those living with hypertension have their blood pressure effectively controlled).

Monte Rosa Therapeutics Announces Positive Results for MRT-8102 in GFORCE-1 Phase 1 Study Showing Normalization of Key Pathogenic Drivers of ASCVD in Subjects with Elevated CVD Risk

In obese subjects at elevated cardiovascular disease (CVD) risk, MRT-8102, a NEK7-directed molecular glue degrader in development for the treatment of NLRP3/IL-1-driven inflammatory diseases, normalized multiple atherosclerotic cardiovascular disease (ASCVD) pathogenic drivers of local plaque inflammation, thrombosis, and systemic inflammation Robust and sustained NEK7 degradation observed across 5 mg, 20 mg, and 40 mg once-daily dose levels, with similar biomarker reductions at all three doses HMGB1, calprotectin, S100A12, and SAA, damage-associated molecular patterns (DAMPs) linked to atherosclerotic plaque progression and instability, and IL-1β, all established key pathogenic drivers of ASCVD, were reduced to levels comparable to normal physiological levels seen in the healthy volunteer population of Phase 1 study MRT-8102 was safe and well tolerated over four weeks of dosing and four weeks of follow-up, with no serious adverse events, treatment-emergent adverse event rates comparable to placebo, and no evidence of increased infection risk Phase 2 trials planned across coronary artery disease (GFORCE-2), gout (GEMINI-1), and moderate to severe hidradenitis suppurativa (GALAXY-1) Company to host conference call and webcast today, Oct. 1, at 8:00 a.m. ET BOSTON, Oct. 01, 2026 (GLOBE NEWSWIRE) — Monte Rosa Therapeutics, Inc. (Nasdaq: GLUE), a clinical-stage biotechnology company developing novel molecular glue degrader (MGD)-based medicines, today announced positive data from GFORCE-1, a Phase 1 study evaluating MRT-8102, a NEK7-directed MGD being developed for the treatment of inflammatory conditions driven by the NEK7/NLRP3 pathway. “The GFORCE-1 data are highly encouraging and show that MRT-8102 does precisely what we designed it to do: potently and selectively degrade NEK7 and reduce levels of upstream and downstream drivers of residual plaque inflammation and thrombotic risk, and it does so with a differentiated and favorable safety profile,” said Markus Warmuth, M.D., Chief Executive Officer of Monte Rosa Therapeutics. “We are particularly excited about our translational approach, including our unique and broad analysis of the NEK7/NLRP3 upstream pathway, that has allowed us to gain unprecedented insights into how inhibition of the pathway through NEK7 degradation leads to normalization of many of the most important drivers of atherosclerotic plaque formation, progression, and rupture. To that end, we observed compelling reductions of HMGB1, calprotectin, S100A12, SAA, and IL-1β. These DAMPs and cytokines are linked to pathogenic biology within atherosclerotic plaques and vessel walls, and they have been identified as independent risk factors of much greater significance than some of the systemic inflammatory markers evaluated in other studies. We have also seen intriguing correlation of these markers with NLRP3 risk alleles, opening up opportunities for a precision medicine approach. The GFORCE-1 study results, alongside a body of genetic, biological, and clinical data, reinforce our rationale for degrading NEK7 to simultaneously dampen multiple key ASCVD disease drivers that can lead to atherosclerosis, plaque progression, and thrombogenesis. We believe targeting NEK7 at the top of the NLRP3 signaling cascade has the potential to provide a degree of clinical impact across NEK7/NLRP3-driven diseases not achievable by targeting individual downstream cytokines.” Filip Janku, M.D., Ph.D., Chief Medical Officer of Monte Rosa Therapeutics, commented, “We believe our results represent the most comprehensive translational data set reported to date for the NEK7/NLRP3 pathway. GFORCE-1 was designed to provide a broad view of the pharmacodynamic activity and safety of MRT-8102, and the profile we observed further bolsters our enthusiasm for continued development in cardiovascular disease and other indications. With dose selection now informed by these data, we plan to initiate GFORCE-2, a focused Phase 2b study in patients with coronary artery disease, in the first half of 2027. In GFORCE-2, our goal is to connect imaging-based coronary artery plaque assessment, such as fat attenuation index (FAI), with NLRP3 genetics and molecular drivers of local disease pathogenesis and systemic inflammation to inform the design of an innovative and efficient Phase 3 study focused on the most disease-modifiable patient population. We also expect to initiate GEMINI-1, a Phase 2 study in gout, in Q4 2026 or Q1 2027, and GALAXY-1, a Phase 2 study in moderate to severe hidradenitis suppurativa, in the first half of 2027, representing additional substantial opportunities associated with this pathway.” GFORCE-1 (clinicaltrials.gov identifier NCT07119125) enrolled 108 obese subjects with elevated CVD risk, randomized to MRT-8102 at once-daily doses of 5 mg, 20 mg, or 40 mg, or to placebo. The GFORCE-1 study followed best-in-class data from the single ascending dose (SAD) and multiple ascending dose (MAD) cohorts in healthy volunteers, and those data were previously reported. The GFORCE-1 study evaluated safety and tolerability, along with multiple pharmacodynamic markers, over a four-week treatment regimen, followed by four weeks of safety follow-up. Multiple dose levels were explored to support dose selection for further development across various NLRP3-driven diseases, including ASCVD, gout, and hidradenitis suppurativa. Summary of GFORCE-1 Results GFORCE-1 enrolled 108 obese subjects with elevated CVD risk, randomized to MRT-8102 at 5 mg, 20 mg, or 40 mg once daily, or to placebo, for 4 weeks of dosing and 4 weeks of follow-up safety evaluation.MRT-8102 achieved robust and sustained NEK7 degradation, with a median reduction of approximately 80 to 90% observed across all three dose levels.Both local drivers of plaque growth and progression and systemic inflammatory biomarkers were substantially reduced following MRT-8102 administration at all three dose levels to levels comparable with baseline values from Phase 1 healthy volunteers.Median values of calprotectin, S100A12, and SAA, which are critical drivers of ASCVD progression, were lowered by 56%, 46%, and 51%, respectively.Median values of HMGB1 and IL-1β were elevated at baseline and decreased toward healthy volunteer levels following treatment among subjects with elevated levels. Median reductions were 40% and 37%, respectively, in subjects with levels in the top quartile.The thrombogenic factor fibrinogen was reduced by a median of 28% to levels consistent with those in healthy volunteers.IL-6 and hsCRP, biomarkers of systemic inflammation, were reduced by a median of 54% and 85%, respectively. All three doses of MRT-8102 resulted in rapid and sustained reductions in hsCRP to levels comparable to baseline in healthy volunteers from the healthy volunteer cohorts.MRT-8102 administration reduced median lipoprotein(a) by 24%, comparable to PCSK9 inhibitors. Unlike IL-6(R) blockade, MRT-8102 treatment did not increase LDL-C or triglyceride levels.Significant increases in baseline biomarker levels were identified in carriers of NLRP3 risk alleles (approximately 30% of the GFORCE-1 population), particularly for DAMPs and IL-1β, suggesting a higher-risk, genetically enrichable population for potential future development.MRT-8102 was well tolerated over four weeks of dosing and four weeks of follow-up. No serious adverse events (SAEs) were reported. Rates of treatment-emergent adverse events (TEAEs) were similar between MRT-8102 and placebo, occurring in 33% of MRT-8102 participants and 30% of placebo participants, with no evidence of increased infection risk. Phase 2 Program for MRT-8102 Monte Rosa expects to initiate multiple Phase 2 studies of MRT-8102 in indications with high unmet need and strong biologic rationale for targeting the NEK7/NLRP3 pathway: GFORCE-2, a focused Phase 2b study of MRT-8102 in patients with stable coronary artery disease and residual inflammation, is expected to initiate in H1 2027. The study design, pending regulatory feedback, will evaluate changes in local pathogenic drivers of ASCVD, systemic biomarkers, and cardiovascular imaging over a six-month dosing period to specifically observe modulation of plaque biology, as well as changes in inflammatory, thrombogenic, and lipid mediators of disease. The Company is working to complete long-term preclinical toxicology studies to support this 6-month study.GEMINI-1, a Phase 2 study of MRT-8102 in patients with gout, is expected to initiate in Q4 2026 or Q1 2027. The study will investigate prevention of flare recurrence following management of acute flares with MRT-8102. Initial data are expected in H2 2027.GALAXY-1, a Phase 2 study of MRT-8102 in patients with moderate to severe hidradenitis suppurativa, is expected to initiate in H1 2027. Investor Conference Call Monte Rosa will host a conference call and webcast presentation today, October 1, 2026, at 8:00 a.m. ET. A webcast of the presentation will be accessible via the “Events & Presentations” section of Monte Rosa’s website at ir.monterosatx.com. Registration for the conference call is available at the following link. An archived version of the webcast will be made available for 30 days following the presentation. About MRT-8102 MRT-8102 is a potent, highly selective, and orally bioavailable investigational molecular glue degrader (MGD) that targets NEK7 for the treatment of inflammatory diseases linked to NLRP3 and IL-1 dysregulation. NEK7 has been shown to be required for NLRP3 inflammasome assembly, activation, and IL-1β release both in vitro and in vivo. NLRP3 inflammasome activation causes pyroptotic cell death, leading to the release of IL-1α/β, IL-18, and DAMPs such as HMGB1, calprotectin, S100A12, and SAA. IL-1β and IL-18 are highly inflammatory cytokines, and DAMPs further stimulate NLRP3 activity while promoting NLRP3-independent cytokine and chemokine production in neighboring cells. Together, DAMPs and IL-1 are responsible for the majority of the local inflammation and tissue changes characteristic of ASCVD, gout, hidradenitis suppurativa, and other indications. The NEK7/NLRP3 inflammasome plays a multifaceted role in plaque biology, contributing to plaque formation, plaque growth, vessel wall alterations, and thrombosis. About Monte Rosa Monte Rosa Therapeutics is a clinical-stage biotechnology company developing highly selective molecular glue degrader (MGD) medicines for patients living with serious diseases. MGDs are small molecule protein degraders that have the potential to treat many diseases that other modalities, including other degraders, cannot. Monte Rosa’s QuEEN™ (Quantitative and Engineered Elimination of Neosubstrates) discovery engine combines AI-guided chemistry, diverse chemical libraries, structural biology, and proteomics to rationally design MGDs with unprecedented selectivity. Monte Rosa has developed the industry’s leading pipeline of first-in-class and only-in-class MGDs, spanning autoimmune and inflammatory diseases, oncology, and beyond, with three programs in the clinic. Monte Rosa has ongoing collaborations with leading pharmaceutical companies in the areas of immunology, oncology, and neurology. For more information, visit www.monterosatx.com. Forward-Looking Statements This communication includes express and implied “forward-looking statements,” including forward-looking statements within the meaning of the Private Securities Litigation Reform Act of 1995. Forward-looking statements include all statements that are not historical facts and in some cases, can be identified by terms such as “may,” “might,” “will,” “could,” “would,” “should,” “expect,” “intend,” “plan,” “objective,” “anticipate,” “believe,” “estimate,” “predict,” “potential,” “continue,” “ongoing,” or the negative of these terms, or other comparable terminology intended to identify statements about the future. Forward-looking statements contained herein include, but are not limited to, statements about our ability to grow our product pipeline, our ability to successfully complete research and further development and commercialization of our drug candidates in current or future indications, including the timing and results of our clinical trials and our ability to conduct and complete clinical trials, statements regarding our progress and speed of development of only-in-class and first-in-class molecular glue degrader therapeutics, statements around the Company’s QuEEN™ discovery engine and the broad potential applications of the platform and the Company’s ability to create long-term value through focused pipeline execution and strategic collaborations, statements about the advancement and timeline of our preclinical and clinical programs, pipeline and the various products therein, statements regarding the positive Phase 1 data of MRT-8102 from the GFORCE-1 study, including the safety, tolerability, target engagement, and other results observed, statements regarding the potential of MRT-8102 to normalize local and systemic pathologic drivers of ASCVD and to address NLRP3/IL-1-driven inflammatory diseases, our expectations regarding the potential of targeting NEK7 at the top of the NLRP3 inflammasome signaling cascade to provide differentiated clinical impact across NLRP3-driven diseases, including compared with targeting individual downstream cytokines, statements regarding the planned initiation of GFORCE-2, a Phase 2b study of MRT-8102 in patients with stable coronary artery disease and residual inflammation, including the expected timing, study design, regulatory feedback regarding the study design, use of coronary artery imaging, NLRP3 genetics and biomarkers to identify a disease-modifiable patient population and inform the design of a potential Phase 3 study, and the completion of long-term preclinical toxicology studies to support GFORCE-2, statements regarding the planned initiation of GEMINI-1, a Phase 2 study of MRT-8102 in patients with gout, including the expected timing, study design and objective of evaluating prevention of flare recurrence, and the expected timing of initial data, statements regarding the planned initiation of GALAXY-1, a Phase 2 study of MRT-8102 in patients with moderate to severe hidradenitis suppurativa, including the expected timing, statements regarding dose selection informed by GFORCE-1 data for further development across various NLRP3-driven diseases, statements regarding a genetically enrichable higher-risk population identified through NLRP3 risk alleles for potential future development, statements regarding opportunities for MRT-8102 in inflammatory diseases and statements around our expectations of success for our programs, among others. By their nature, these statements are subject to numerous risks and uncertainties, including those risks and uncertainties set forth in our most recent Annual Report on Form 10-K for the year ended December 31, 2025, filed with the U.S. Securities and Exchange Commission on March 17, 2026, and any subsequent filings, that could cause actual results, performance or achievement to differ materially and adversely from those anticipated or implied in the statements, as well as the risk that outcomes of preclinical studies may not be predictive of clinical trial results and the risk that initial or interim results from a clinical trial may not be predictive of the final results of the trial or the results of future trials. You should not rely upon forward-looking statements as predictions of future events. Although our management believes that the expectations reflected in our statements are reasonable, we cannot guarantee that the future results, performance, or events and circumstances described in the forward-looking statements will be achieved or occur. Recipients are cautioned not to place undue reliance on these forward-looking statements, which speak only as of the date such statements are made and should not be construed as statements of fact. We undertake no obligation to publicly update any forward-looking statements, whether as a result of new information, any future presentations, or otherwise, except as required by applicable law. Certain information contained in these materials and any statements made orally during any presentation of these materials that relate to the materials or are based on studies, publications, surveys and other data obtained from third-party sources and our own internal estimates and research. While we believe these third-party studies, publications, surveys and other data to be reliable as of the date of these materials, we have not independently verified, and make no representations as to the adequacy, fairness, accuracy or completeness of, any information obtained from third-party sources. In addition, no independent source has evaluated the reasonableness or accuracy of our internal estimates or research and no reliance should be made on any information or statements made in these materials relating to or based on such internal estimates and research. InvestorsAndrew Funderburkir@monterosatx.com MediaCory Tromblee, Scient PRmedia@monterosatx.com

Kestra Medical Technologies Earns Five-Star Rating on Newsweek’s List of America’s Greatest Mid-Market Companies

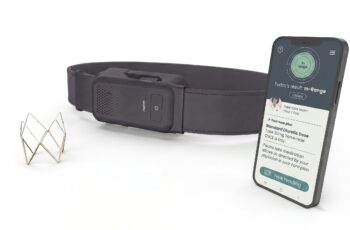
KIRKLAND, Wash., Oct. 01, 2026 (GLOBE NEWSWIRE) — Kestra Medical Technologies, Ltd. (Nasdaq: KMTS), a leading wearable medical device and digital healthcare company, has earned a five-star rating on Newsweek’s list of America’s Greatest Mid-Market Companies 2026. Only 30 of the 100 honorees earned five stars, and Kestra was the only company in the Medical Equipment Manufacturing category to receive that rating. “In less than a decade, our team brought the ASSURE® WCD to market and built a rapidly growing commercial business, giving clinicians a differentiated choice for protecting patients during periods of cardiac risk and recovery,” said Brian Webster, President and Chief Executive Officer of Kestra Medical Technologies. “This five-star rating reflects the strength of the Kestra team and the company we’ve built. We’re now positioned to extend our reach, advance our Cardiac Recovery System® platform, and pursue a bigger vision for wearable cardiac protection.” For its inaugural ranking, Newsweek and Plant-A Insights Group assessed publicly traded U.S. mid-market companies across financial performance, workforce experience, innovation, and sustainability. Their analysis drew on business metrics, third-party workforce data, and more than 2.7 million company reviews. “Newsweek’s ranking of America’s Greatest Mid-Market Companies recognizes businesses that are proving that meaningful growth and lasting impact aren’t limited to the largest corporations,” added Ryan Kinney, Senior Vice President, Newsweek. “The ranking shines a spotlight on midsize companies that are strengthening their industries, creating opportunities for team members and customers, and demonstrating the ambition and resilience needed to succeed in a competitive marketplace.” About KestraKestra Medical Technologies, Ltd. is a commercial-stage wearable medical device and digital healthcare company focused on transforming patient outcomes in cardiovascular disease using monitoring and therapeutic intervention technologies that are intuitive, intelligent, and connected. For more information, please visit www.kestramedical.com.  Forward-Looking StatementsThis press release contains forward-looking statements that involve risks and uncertainties. Actual results may differ materially from those expressed or implied by these statements. For a discussion of factors that could cause actual results to differ, see Kestra’s filings with the U.S. Securities and Exchange Commission. Kestra undertakes no obligation to update any forward-looking statements except as required by law. CONTACT: Media contact 
Rhiannon Pickus 
rhiannon.pickus@kestramedical.com 
Eli lilly and company logo

Lilly’s oral GLP-1, Foundayo (orforglipron), demonstrated cardiovascular safety alongside sustained A1C reduction and weight loss in its largest and longest type 2 diabetes study

In ACHIEVE-4, participants taking Foundayo experienced a 16% lower risk of MACE-4 and a 23% lower risk of MACE-3, meeting the primary objective for non-inferiority compared to insulin glargine In pre-planned analyses, Foundayo showed a 53% lower risk of cardiovascular death and a 57%…